PK terminology • Exposure interpretation

Voriconazole–Cyclosporine Interaction Terminology & PK Interpretation

Cyclosporine-interaction terminology describes pharmacokinetic relationships in a documentation context rather than clinical instruction. In voriconazole–cyclosporine discussions, interaction language can distinguish systemic input, bioavailability, absorption variability, distribution, metabolic capacity, and clearance. Formulation is also relevant because a tablet, oral suspension, and IV form represent different administration contexts and input characteristics that can influence interpretation of observed exposure. Inhibitor terminology is descriptive: it identifies a mechanistic relationship in which one substance alters enzyme-mediated disposition of another, without implying a management action. Interaction-related PK variability can therefore be expressed through changes in exposure metrics, concentration-time relationships, or apparent clearance rather than through therapeutic conclusions. These terms provide a structured vocabulary for describing pharmacokinetic observations while keeping mechanistic interpretation separate from clinical decision-making.

Interpretation of voriconazole–cyclosporine interaction terminology also incorporates metabolism, CYP2C19 phenotype, nonlinear kinetics, and clearance. CYP2C19 phenotype terminology can describe genetically associated differences in voriconazole metabolic capacity, while nonlinear kinetics terminology describes concentration-dependent or otherwise disproportionate changes in pharmacokinetic parameters. These concepts may contribute to between-subject and within-subject variability in systemic exposure and can affect how cyclosporine concentrations are represented in pharmacokinetic documentation. The terms do not themselves establish a clinical outcome. Instead, they provide a vocabulary for separating formulation-dependent input, systemic disposition, enzyme-mediated interaction mechanisms, and measured exposure from clinical interpretation. This distinction is useful when several PK determinants operate simultaneously and a concentration change cannot reasonably be attributed to one isolated process.

Temporal descriptors provide another layer of neutral interpretation. Tmax and Cmax describe the timing and magnitude of a concentration peak, while half-life describes the temporal decline associated with drug disposition. TDM terminology can describe concentration measurement and longitudinal observation as documentation concepts without specifying clinical actions or target concentrations. In a voriconazole–cyclosporine interaction record, these descriptors can be considered alongside bioavailability, absorption variability, distribution, metabolism, and clearance to characterize observed concentration-time behavior. The resulting framework is pharmacokinetic rather than therapeutic: it explains terminology used to describe systemic input, interaction mechanisms, exposure variability, and temporal concentration patterns. Formulation, metabolic phenotype, sampling schedule, assay characteristics, and kinetic assumptions can all be documented as contextual variables without introducing dose-adjustment guidance or interaction-management recommendations.

Cyclosporine-Interaction Terminology Foundations
Cyclosporine levels

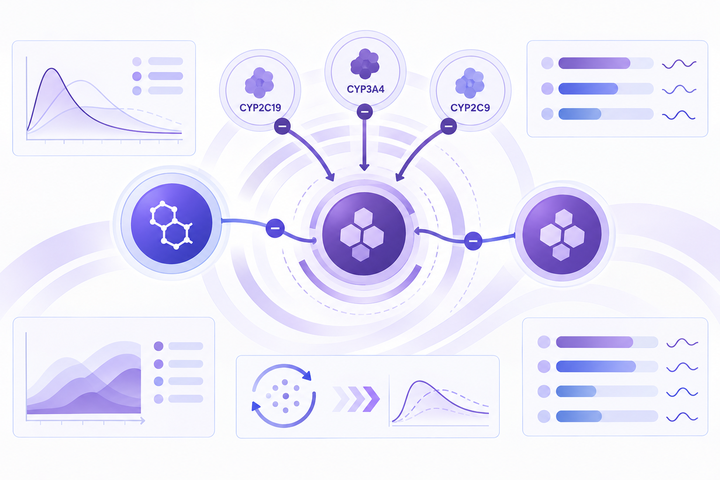
Cyclosporine-interaction terminology begins with the distinction between a perpetrator and a victim drug. In pharmacokinetic interaction documentation, voriconazole may be characterized as the interacting substance and cyclosporine as the affected substrate when the record focuses on altered disposition. Terms such as substrate, inhibitor, enzyme-mediated interaction, exposure change, systemic concentration, and apparent clearance describe mechanistic relationships. The term interaction does not by itself specify whether an observed concentration difference has clinical significance. Instead, it identifies a relationship requiring pharmacokinetic characterization. Documentation can separately describe baseline exposure, co-exposure, post-interaction exposure, concentration-time profiles, and variability. This separation prevents mechanistic terminology from being interpreted as treatment instruction. A documented exposure ratio, for example, is a quantitative descriptor, whereas an inhibitor designation is a mechanistic interpretation. Keeping these categories distinct improves terminology precision and preserves the difference between measured pharmacokinetic data and clinical conclusions.

Formulation and route provide another foundation for interpretation. A tablet or oral suspension introduces an absorption phase, whereas an IV form represents systemic input without conventional gastrointestinal absorption. Terms such as bioavailability, absorption rate, absorption extent, and absorption variability therefore describe components that can influence observed exposure independently of metabolic interaction. Distribution can subsequently influence the relationship between plasma concentration and tissue concentration. These distinctions matter because a measured concentration difference can reflect multiple pharmacokinetic determinants rather than one interaction mechanism. Documentation may therefore specify formulation, route, sampling conditions, and relevant exposure metrics before interpreting a concentration-time pattern. Such terminology allows systemic input to be considered separately from metabolism, clearance, and other disposition processes.

Interaction terminology can then be connected to quantitative PK descriptors. Exposure may be represented by area-under-the-curve measures, maximum concentration, trough concentration, average concentration, or concentration-time profiles. Apparent clearance expresses elimination relative to systemic exposure, while half-life describes the temporal persistence of concentration under a defined kinetic model. A change in one descriptor does not automatically establish a particular mechanism because several parameters can change simultaneously. Documentation therefore benefits from separating input, absorption, distribution, metabolism, elimination, and measurement variability. This vocabulary supports a mechanistic description of voriconazole–cyclosporine relationships while avoiding dose-adjustment language, therapeutic interpretation, risk classification, or interaction-management recommendations.

Interaction Term Mechanistic Basis Exposure Role
Perpetrator Substance producing a pharmacokinetic effect on another drug Frames the source of an observed disposition change
Victim drug Drug whose pharmacokinetics are altered in an interaction description Provides the exposure variable being characterized
Substrate Drug undergoing enzyme-mediated or transporter-associated disposition Connects metabolic processes with systemic exposure
Inhibitor Substance associated with reduced activity of a metabolic pathway Can be described through altered exposure or apparent clearance
Interaction effect Observed pharmacokinetic difference associated with co-exposure Describes changes in concentration-time or exposure metrics

Inhibitor Terminology (Mechanistic)

Inhibitor terminology describes how one substance affects enzyme-mediated metabolism of another substance. A reversible inhibitor can be characterized according to the persistence and concentration dependence of its inhibitory effect, whereas time-dependent inhibition describes an effect whose magnitude may depend on exposure duration or enzyme turnover. Mechanistic inhibition terminology can also distinguish competitive, noncompetitive, uncompetitive, and mixed patterns when experimental evidence supports those classifications. In a voriconazole–cyclosporine context, such terminology is used to characterize potential changes in metabolic disposition rather than to prescribe an action. The documentation focus is the relationship between enzyme activity, substrate availability, and observed systemic exposure. Inhibition strength may be described through experimental parameters, but those parameters remain dependent on experimental conditions, substrate concentration, enzyme system, and assay design.

CYP-linked terminology adds pathway specificity. An interaction record may identify an enzyme system, substrate relationship, inhibitor relationship, or phenotype-dependent metabolic characteristic. The relevant language should distinguish direct experimental findings from inferred mechanisms because concentration changes can arise from multiple processes. Inhibitory potency can be described through parameters such as inhibitory concentration or inhibition constants, but these measures do not automatically establish a clinical interpretation. Apparent clearance can provide a quantitative description of altered elimination, while exposure metrics can characterize the resulting concentration-time relationship. The terms remain descriptive even when an exposure difference is reproducible. This distinction is especially important when the interacting drugs have multiple metabolic and transporter pathways.

The temporal dimension of inhibition is also relevant. Rapidly reversible inhibition, delayed inhibition, and recovery after removal of an inhibitor can produce different concentration-time patterns depending on enzyme turnover, drug half-life, exposure history, and sampling design. Documentation may therefore distinguish an interaction mechanism from the observed magnitude and timing of an exposure change. When formulation, metabolic phenotype, nonlinear processes, or multiple pathways are present, the observed concentration profile may reflect several concurrent determinants. Inhibitor terminology should consequently be treated as a mechanistic vocabulary for pharmacokinetic description, not as shorthand for treatment selection, dose modification, risk categorization, safety interpretation, or interaction-management guidance.

Inhibitor Term Mechanistic Link PK Interpretation
Reversible inhibition Inhibitory effect can diminish as inhibitor concentration changes Exposure may vary with concentration and timing
Competitive inhibition Inhibitor and substrate interact with a shared enzyme process Provides a model for altered substrate metabolism
Time-dependent inhibition Inhibitory magnitude changes with exposure duration or enzyme turnover Can produce delayed or persistent PK effects
Inhibition constant Quantitative descriptor of inhibitor–enzyme interaction under defined conditions Supports mechanistic characterization rather than clinical interpretation
Pathway inhibition Reduced activity of a specified metabolic pathway May be reflected in altered exposure or clearance descriptors

Systemic Exposure Variability

Systemic exposure variability describes differences in drug concentration or exposure between observations, individuals, formulations, or study periods. In voriconazole–cyclosporine documentation, variability can arise before systemic circulation through formulation characteristics, absorption rate, absorption extent, and bioavailability. Once systemic input occurs, distribution, metabolic activity, transporter processes, and elimination contribute additional variability. Interindividual variability refers to differences between people, whereas intraindividual variability describes changes within the same person across observations. Residual variability can reflect assay variation, sampling uncertainty, model misspecification, or unmeasured biological factors. These categories help distinguish sources of dispersion without assigning a clinical meaning to observed values. Documentation may report variability using descriptive statistics, concentration ratios, model-derived random effects, or other quantitative measures, depending on study design.

Bioavailability terminology is especially important for oral formulations because systemic exposure depends on the fraction of administered drug reaching circulation and the rate at which that input occurs. Tablet and oral-suspension formulations can differ in disintegration, dissolution, gastrointestinal transit, and absorption characteristics. An IV formulation changes the input framework because systemic availability is not limited by gastrointestinal absorption. Consequently, an interaction comparison across formulations should distinguish changes attributable to systemic input from changes attributable to metabolic disposition. Absorption variability may alter Tmax and Cmax while leaving other exposure descriptors differently affected, depending on the underlying process, formulation, and sampling schedule. These distinctions help prevent input-related variation from being automatically attributed to enzyme-mediated interaction.

Systemic exposure can be represented using area-under-the-curve measures, maximum concentration, minimum or trough concentration, average concentration, and concentration-time profiles. Exposure variability may therefore be expressed as absolute differences, ratios, coefficients of variation, or model-derived parameters. Such metrics describe the distribution of pharmacokinetic observations but do not independently establish mechanism. When voriconazole and cyclosporine are documented together, interpretation may require consideration of formulation, bioavailability, absorption variability, enzyme-mediated metabolism, clearance, sampling schedule, assay characteristics, and underlying kinetic assumptions. The resulting terminology provides a structured description of exposure behavior while avoiding safety claims, therapeutic thresholds, risk classification, or management recommendations.

Exposure Variable Mechanistic Basis Cyclosporine-Context Role
AUC Integrated concentration over a defined time interval Describes systemic exposure across time
Cmax Maximum observed or modeled concentration Characterizes peak exposure behavior
Trough concentration Concentration measured near a defined dosing interval point Provides a time-specific exposure observation
Bioavailability Fraction and rate of systemic input after administration Separates input characteristics from disposition
Interindividual variability Differences in PK parameters between individuals Describes population-level exposure dispersion

Metabolism, CYP2C19 & Nonlinear Kinetics

Metabolism terminology describes biochemical transformation that contributes to drug disposition. In voriconazole documentation, CYP2C19 is a relevant pharmacogenetic descriptor because phenotype terminology can distinguish genetically associated differences in metabolic capacity. Terms such as poor, intermediate, normal, rapid, or ultrarapid metabolizer describe phenotype categories under defined classification systems, although observed pharmacokinetic phenotype can also reflect environmental, physiological, concomitant-drug, and formulation factors. A CYP2C19 phenotype therefore represents one determinant within a larger metabolic system. When discussing cyclosporine interaction PK, the distinction between voriconazole metabolic phenotype and cyclosporine disposition pathways is important because each compound can have different metabolic determinants and interaction mechanisms. Documentation can identify these variables separately rather than treating CYP2C19 phenotype as a universal descriptor of both drugs.

Nonlinear kinetics describes situations in which pharmacokinetic parameters or exposure do not change proportionally with input or concentration. Nonlinearity can result from saturable metabolism, saturable transport, concentration-dependent binding, capacity-limited elimination, or other processes. In documentation, nonlinear behavior can complicate direct comparison of exposure across different systemic inputs because an observed concentration change may not scale proportionally with an input change. The term does not identify a single mechanism unless supporting evidence is available. Modeling may distinguish linear and nonlinear components, estimate concentration-dependent parameters, and evaluate whether apparent clearance changes with concentration or exposure. Such analyses remain descriptive and depend on the data structure, sampling design, and selected model.

CYP-linked interaction terminology should distinguish enzyme inhibition from enzyme induction, although their mechanisms and temporal behavior differ. Inhibitor terminology describes reduced activity of a metabolic pathway, whereas induction terminology describes increased expression or activity over an appropriate biological timescale. The resulting effects on substrate exposure depend on pathway contribution, competing pathways, metabolic capacity, and timing. For voriconazole–cyclosporine documentation, CYP2C19 phenotype, metabolic capacity, enzyme inhibition, nonlinear kinetics, and apparent clearance can be recorded as separate explanatory variables. This preserves mechanistic precision without converting pharmacokinetic terminology into therapeutic recommendations, risk classifications, or interaction-management decisions.

Metabolic Factor CYP Connection Exposure Impact
CYP2C19 phenotype Genetically associated variation in CYP2C19 activity Can contribute to variability in voriconazole disposition
Enzyme inhibition Reduced activity of a metabolic pathway May alter substrate clearance and systemic exposure
Enzyme induction Increased expression or activity of a metabolic pathway May alter substrate disposition over time
Nonlinear elimination Pathway capacity or concentration dependence affects disposition Exposure may not scale proportionally with input
Metabolic capacity Combined functional ability of relevant pathways to transform drug Contributes to between-subject and within-subject variability

Distribution, Clearance & Temporal PK Descriptors

Distribution describes movement of drug between systemic circulation and tissues and is commonly represented through apparent volume of distribution, tissue partitioning, plasma protein binding, and related parameters. In interaction documentation, distribution terminology helps separate changes in concentration from changes in total systemic disposition. Clearance describes the hypothetical volume of plasma or blood from which drug is completely removed per unit time and can be expressed as systemic, hepatic, renal, or apparent clearance depending on the analytical framework. When an interaction changes metabolic disposition, an observed alteration in apparent clearance can provide a quantitative descriptor, but it does not alone identify every contributing mechanism. Other factors, including bioavailability and distribution, can affect how clearance is estimated or interpreted.

Temporal descriptors provide a complementary view of interaction-related PK. Tmax and Cmax describe the time to peak concentration and the peak magnitude, respectively. These descriptors are sensitive to absorption rate, formulation, sampling frequency, and systemic disposition. Half-life describes the time required for concentration to decline by a specified fraction under the relevant kinetic model and can be influenced by clearance and apparent distribution volume. A concentration-time profile therefore contains information that cannot always be captured by a single exposure metric. Documentation may use multiple descriptors to distinguish input-related changes from elimination-related changes, while recognizing that temporal parameters are dependent on the sampling design and model assumptions.

Longitudinal concentration measurement can be described with TDM terminology when documentation concerns repeated drug-concentration observations. TDM is a measurement framework rather than, by itself, a statement about treatment action. Sampling time, assay method, concentration matrix, formulation, exposure history, and elapsed time from administration can all influence interpretation of a measured concentration. For voriconazole–cyclosporine interaction records, these variables can be documented alongside distribution, clearance, Tmax, Cmax, and half-life. Such descriptors support characterization of temporal PK behavior while maintaining a neutral distinction between measured pharmacokinetic observations and clinical decision-making. The same terminology can be applied to observed, modeled, population, and individual concentration profiles when the analytical context is explicitly stated.

PK Descriptor Mechanistic Connection Documentation Context
Distribution Movement between systemic circulation and tissues Describes concentration partitioning and apparent volume relationships
Clearance Systemic elimination relative to circulating drug Quantifies disposition without independently assigning mechanism
Tmax Timing of maximum observed or modeled concentration Characterizes temporal peak behavior
Cmax Magnitude of maximum observed or modeled concentration Characterizes peak exposure
Half-life Temporal decline determined by disposition parameters Describes persistence of concentration under the relevant kinetic model

Documentation Interpretation Factors (neutral)

Pharmacokinetic documentation should distinguish observed findings from mechanistic interpretation. A concentration difference is an observation; an exposure ratio is a quantitative comparison; an inhibitor designation is a mechanistic interpretation; and a formulation designation identifies the systemic-input context. These categories should not be conflated. Relevant documentation factors include formulation, route, sampling schedule, assay methodology, exposure history, metabolic phenotype, physiological variables when available, and model assumptions. The presence of several simultaneous variables can create uncertainty about which mechanism accounts for a specific exposure pattern. Neutral terminology therefore emphasizes what was measured, under what conditions, and which mechanistic explanation is supported. A documented association between co-exposure and concentration change does not automatically establish causality.

Formulation-dependent terminology is particularly important when comparing oral and intravenous exposure. A tablet or oral suspension introduces absorption processes that can affect bioavailability, absorption rate, Tmax, and Cmax. An IV form provides a different systemic-input profile and therefore changes the interpretation of apparent bioavailability and absorption-related variability. If interaction observations are compared across formulations, documentation can identify whether a difference concerns input, systemic disposition, or both. Similarly, repeated concentration measurements should be associated with their sampling times because temporal descriptors can otherwise be difficult to interpret. These distinctions help maintain consistency when documenting interaction-related PK variability and prevent formulation effects from being automatically attributed to metabolic inhibition.

Uncertainty can arise from incomplete sampling, sparse concentration data, assay variability, unobserved covariates, phenotype heterogeneity, nonlinear kinetics, and differences between population-level and individual-level parameters. Model-derived estimates may therefore differ from directly observed measurements, and apparent changes in clearance or exposure may depend on the analytical framework. Documentation can explicitly label assumptions, data limitations, and competing mechanistic explanations rather than presenting an inferred mechanism as established fact. This approach keeps voriconazole–cyclosporine interaction terminology focused on pharmacokinetic description, systemic exposure, and evidence-supported interpretation. It avoids introducing safety conclusions, therapeutic thresholds, dose-adjustment instructions, risk stratification, or interaction-management recommendations into a terminology-focused record.

Interpretation Factor Mechanistic Basis Documentation Role
Formulation Changes route, systemic input, dissolution, or absorption characteristics Defines the PK context of an interaction observation
Sampling schedule Determines which portions of the concentration-time profile are observed Constrains temporal and exposure interpretation
Assay variability Analytical measurement uncertainty Provides context for observed concentration dispersion
Phenotype variability Differences in genetically or physiologically influenced metabolic capacity Documents a source of between-subject PK variation
Model assumptions Structural and statistical choices used to estimate PK parameters Clarifies how inferred parameters were derived

Frequently Asked Questions

Cyclosporine-interaction terminology describes pharmacokinetic relationships between cyclosporine and another substance, including systemic input, metabolism, inhibition, clearance, concentration, and exposure. The terminology is mechanistic and documentation-focused. It can identify how an interaction is characterized through concentration-time data or PK parameters without implying a therapeutic decision, dose change, safety conclusion, risk classification, or specific interaction-management action.

Inhibitor terminology describes a substance that reduces activity of a metabolic enzyme or pathway under defined experimental or pharmacokinetic conditions. Terms such as reversible, competitive, or time-dependent inhibition describe different mechanistic patterns. In a cyclosporine interaction record, inhibitor language identifies a potential disposition mechanism rather than providing instructions about treatment, dosing, monitoring actions, risk classification, or clinical management.

Systemic exposure can vary because of differences in bioavailability, absorption, formulation, metabolic capacity, enzyme activity, distribution, clearance, sampling conditions, and assay characteristics. Between-subject and within-subject variability may both contribute. During interaction documentation, these factors are considered potential contributors to observed concentration differences, without assuming that any single factor completely explains the measured pharmacokinetic pattern or establishes clinical significance.

Hepatic metabolism can be described through enzyme-mediated biotransformation, metabolic capacity, pathway contribution, inhibition, induction, and apparent clearance. Documentation may distinguish the metabolic pathway involved from the measured concentration or exposure effect. This distinction is important because an observed PK change can have multiple determinants. Hepatic metabolism terminology therefore describes disposition mechanisms without establishing therapeutic significance, safety conclusions, or clinical action.

Nonlinear kinetics means that pharmacokinetic parameters or exposure do not change proportionally with dose, concentration, or systemic input under the relevant conditions. Potential mechanisms include saturable metabolism, transport, binding, or elimination. In interaction documentation, nonlinear terminology signals that simple proportional assumptions may not adequately describe observed exposure. It does not independently identify a specific mechanism or imply a clinical recommendation.

Temporal PK descriptors include Tmax, Cmax, concentration-time profiles, trough concentrations, and half-life. Tmax identifies the timing of a peak, Cmax describes peak magnitude, and half-life characterizes concentration decline under the applicable kinetic model. These descriptors can help distinguish input-related and disposition-related patterns. Their interpretation depends on sampling times, formulation, assay methods, exposure history, and the underlying pharmacokinetic model.

Documentation uncertainty can result from sparse sampling, incomplete concentration-time profiles, assay variability, formulation changes, phenotype heterogeneity, nonlinear kinetics, unmeasured covariates, and differences between observed and model-derived parameters. A measured exposure difference may therefore support several possible interpretations. Neutral PK documentation can distinguish observations, quantitative estimates, assumptions, and mechanistic hypotheses rather than presenting an uncertain explanation as established fact.

Formulation-dependent terminology identifies how systemic input differs between tablets, oral suspensions, and intravenous formulations. Oral formulations involve absorption and bioavailability, whereas intravenous administration provides systemic input without conventional gastrointestinal absorption. These differences can influence concentration-time descriptors and exposure interpretation. Consequently, an interaction observation should be interpreted in relation to its formulation and route rather than treating all administration contexts as pharmacokinetically identical.

Mayo Clinic — Voriconazole Overview EMA — Voriconazole (VFEND) EPAR MedlinePlus — Voriconazole Drugs.com — Voriconazole Monograph PubMed — Voriconazole Studies